Note
Go to the end to download the full example code.
Exact diagonalization
Here we show how to find the eigenvalues and eigenvectors of a many-body Hamiltonian of fermions with Coulomb interactions. We then determine their spin and orbital angular momentum and how this changes when we switch on spin-orbit coupling.
Import the necessary modules.
import numpy as np
from math import comb
import matplotlib.pyplot as plt
import edrixs
Parameters
Define the orbital angular momentum number \(l=1\) (i.e. a p shell), the number of spin-orbitals, the occupancy and the Slater integrals. \(F^{k}\) with \(k=0,2\):
Coulomb interactions
The Coulomb interactions in EDRIXS are described by a tensor. Understanding this in full is complicated and requires careful consideration of the symmetry of the interactions. See Coulomb interactions for more discussion if desired. EDRIXS can construct the matrix via
Create basis
Now we build the binary form of the Fock basis \(|F\rangle\) (we consider it preferable to use the standard \(F\) and trust the reader to avoid confusing it with the interaction parameters.) The Fock basis is the simplest legitimate form for the basis and it consists of a series of 1s and 0s where 1 means occupied and 0 means empty. These are in order up, down, up, down, up, down. For computational efficiency, these are encoded as integers, with orbital zero in the least significant bit. We print occupations in orbital order.
basis = edrixs.get_fock_basis_int(norb, noccu)
print("Integer\tOccupations (orbital 0 first)")
for state in basis.basis_int:
    occupations = "".join(str((state >> orbital) & 1) for orbital in range(norb))
    print(f"{state:2d}\t{occupations}")
Integer Occupations (orbital 0 first)
 3      110000
 5      101000
 9      100100
17      100010
33      100001
 6      011000
10      010100
18      010010
34      010001
12      001100
20      001010
36      001001
24      000110
40      000101
48      000011
We expect the number of these states to be given by the mathematical combination of two electrons distributed among six states (three spin-orbitals with two spins per orbital).
We predict C(norb=6, noccu=2)=15 states and we got 15, which is reassuring!
Note that in more complicated problems with both valence and core electrons, the edrixs convention is to list the valence electrons first.
Transform interactions into Fock basis
edrixs works by initially creating a Hamiltonian matrix
\(\hat{H}\) in the single particle basis and then transforming into
our chosen Fock basis. The Hamiltonian contains a two fermion term built from
the single particle matrix \(t_{ij}\) (passed as emat)
\[\hat{H}_{2} = \langle F_l|\sum_{ij}t_{ij}\hat{f}_{i}^{\dagger} \hat{f}_{j}|F_r\rangle\]
and a four fermion term built from the interaction tensor \(U_{ijkl}\)
(passed as umat)
\[\hat{H}_{4} = \langle F_l|\sum_{ijkl}U_{ijkl}\hat{f}_{i}^{\dagger} \hat{f}_{j}^{\dagger} \hat{f}_{k}\hat{f}_{l}|F_r\rangle\]
For now we just build the umat piece. backend='dense'
tells edrixs to build the full matrix including the zeros.
Diagonalize the matrix
Diagonalization returns eigenvalues
e and eigenvectors v where eigenvalue e[i] corresponds
to eigenvector v[:,i].
15 eigenvalues and 15 eigenvectors 15 elements long.
Computing expectation values
To interpret the results, it is informative to compute the expectation values related to the spin \(\mathbf{S}\), orbital \(\mathbf{L}\), and total \(\mathbf{J}\) angular momentum. We first load the relevant matrices for these quantities for a p atomic shell. We need to specify that we would like to include spin when loading the orbital operator.
We again transform these matrices to our Fock basis to build the operators.
Each is a one-body operator, so it is passed as the emat argument with
umat=None. They have three Cartesian components, which we build one at
a time.
Recall that quantum mechanics forbids us from knowing all three Cartesian components of angular momentum at once, so we want to compute the squares of these operators i.e.
\[\begin{split}\mathbf{S}^2 = S^2_x + S^2_y + S^2_z\\ \mathbf{L}^2 = L^2_x + L^2_y + L^2_z\\ \mathbf{J}^2 = J^2_x + J^2_y + J^2_z\end{split}\]
Remember that the eigenvalues of \(\mathbf{S}^2\) are in the form \(S(S+1)\) etc. and that they can be obtained by calculating the projection of the operators onto our eigenvectors.
We can determine the degeneracy of the eigenvalues numerically and print out the values as follows
e = np.round(e, decimals=6)
degeneracy = [sum(eval == e) for eval in e]
header = "{:<3s}\t{:>8s}\t{:>8s}\t{:>8s}\t{:>8s}"
print(header.format("#  ", "E  ", "S(S+1)", "L(L+1)", "Degen."))
for i, eigenvalue in enumerate(e):
    values_list = [i, eigenvalue, S2_val[i], L2_val[i], degeneracy[i]]
    print("{:<3d}\t{:8.3f}\t{:8.3f}\t{:8.3f}\t{:>3d}".format(*values_list))
#            E            S(S+1)          L(L+1)          Degen.
0          3.800           2.000           2.000          9
1          3.800           2.000           2.000          9
2          3.800           2.000           2.000          9
3          3.800           2.000           2.000          9
4          3.800           2.000           2.000          9
5          3.800           2.000           2.000          9
6          3.800           2.000           2.000          9
7          3.800           2.000           2.000          9
8          3.800           2.000           2.000          9
9          4.040           0.000           6.000          5
10         4.040           0.000           6.000          5
11         4.040           0.000           6.000          5
12         4.040           0.000           6.000          5
13         4.040           0.000           6.000          5
14         4.400           0.000          -0.000          1
We see \(S=0\) and \(S=1\) states coming from the two combinations of the spin 1/2 particles. \(L\) can take values of 0, 1, 2. Remember that spin states have degeneracy of \(2S+1\) and the same is true for orbital states. We must multiply these \(S\) and \(L\) degeneracies to get the total degeneracy. Since these particles are fermions, the overall state must be antisymmetric, which dictates the allowed combinations of \(S\) and \(L\).
Energy level diagram
Let us show our findings graphically
fig, ax = plt.subplots()
for i, eigenvalue in enumerate(np.unique(e)):
    art = ax.plot([0, 1], [eigenvalue, eigenvalue], '-',  color='C{}'.format(i))
    ind = np.where(eigenvalue == e)[0][0]
    L = (-1 + np.sqrt(1 + 4*L2_val[ind]))/2
    S = (-1 + np.sqrt(1 + 4*S2_val[ind]))/2
    message = "L={:.0f}, S={:.0f} ({:.0f})"
    ax.text(1, eigenvalue, message.format(L, S, degeneracy[ind]),
            horizontalalignment='right',
            verticalalignment='bottom',
            color='C{}'.format(i))

ax.set_ylabel('Energy')
for loc in ['right', 'top', 'bottom']:
    ax.spines[loc].set_visible(False)

ax.yaxis.set_ticks_position('left')
ax.set_xticks([])
plt.show()
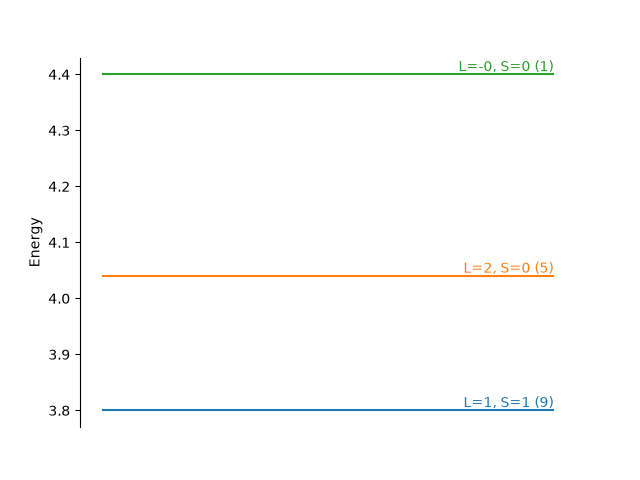
We see Hund’s rules in action! Rule 1 says that the highest spin \(S=1\) state has the lowest energy. Of the two \(S=0\) states, the one with the larger orbital angular momentum (here \(L=2\)) is lower in energy, following rule 2.
Spin orbit coupling
For fun, we can see how this changes when we add spin orbit coupling (SOC). This is a two-fermion operator that we create, transform into the Fock basis and add to the prior Hamiltonian. To make things easy, let us make the SOC small so that the LS coupling approximation is valid and we can still track the states.
Then, we redo the diagonalization and print the results.
e2, v2 = edrixs.ed(H2, num_evals=len(basis))
e2 = np.round(e2, decimals=6)
degeneracy2 = [sum(eval == e2) for eval in e2]
print()
message = "With SOC\n {:<3s}\t{:>8s}\t{:>8s}\t{:>8s}\t{:>8s}\t{:>8s}"
print(message.format("#", "E", "S(S+1)", "L(L+1)", "J(J+1)", "degen."))
J2_val_soc = edrixs.cb_op(J2, v2).diagonal().real
L2_val_soc = edrixs.cb_op(L2, v2).diagonal().real
S2_val_soc = edrixs.cb_op(S2, v2).diagonal().real
for i, eigenvalue in enumerate(e2):
    values_list = [i, eigenvalue, S2_val_soc[i], L2_val_soc[i], J2_val_soc[i],
                   degeneracy2[i]]
    print("{:<3d}\t{:8.3f}\t{:8.3f}\t{:8.3f}\t{:8.3f}\t{:8.3f}".format(*values_list))
With SOC
 #             E          S(S+1)          L(L+1)          J(J+1)          degen.
0          3.673           1.927           1.927          -0.000           1.000
1          3.750           2.000           2.000           2.000           3.000
2          3.750           2.000           2.000           2.000           3.000
3          3.750           2.000           2.000           2.000           3.000
4          3.827           1.802           2.396           6.000           5.000
5          3.827           1.802           2.396           6.000           5.000
6          3.827           1.802           2.396           6.000           5.000
7          3.827           1.802           2.396           6.000           5.000
8          3.827           1.802           2.396           6.000           5.000
9          4.063           0.198           5.604           6.000           5.000
10         4.063           0.198           5.604           6.000           5.000
11         4.063           0.198           5.604           6.000           5.000
12         4.063           0.198           5.604           6.000           5.000
13         4.063           0.198           5.604           6.000           5.000
14         4.427           0.073           0.073          -0.000           1.000
and we make an equivalent energy level diagram.
fig, ax = plt.subplots()
for i, eigenvalue in enumerate(np.unique(e2)):
    art = ax.plot([0, 1], [eigenvalue, eigenvalue], '-',  color='C{}'.format(i))
    ind = np.where(eigenvalue == e2)[0][0]
    J = (-1 + np.sqrt(1+4*J2_val_soc[ind]))/2
    message = "J={:.0f} ({:.0f})"
    ax.text(1, eigenvalue, message.format(J, degeneracy2[ind]),
            horizontalalignment='right',
            verticalalignment='bottom',
            color='C{}'.format(i))

ax.set_ylabel('Energy')
for loc in ['right', 'top', 'bottom']:
    ax.spines[loc].set_visible(False)

ax.yaxis.set_ticks_position('left')
ax.set_xticks([])
plt.show()
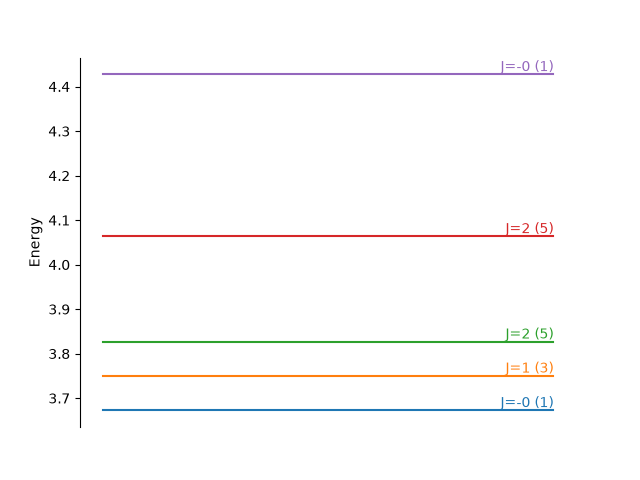
It is clear that we have split the \(S=1\) state, which branches into three states from \(J=|L-S|, |L-S|+1, ..., |L+S|\). Since the shell is less than half full, Hund’s third rule dictates that the smaller \(J\) states have the lower energies.
Total running time of the script: (0 minutes 2.080 seconds)